Insights

The Camelid Ceiling

Why animal sequence, minority VHH output, and constrained diversity move the burden downstream

Camelid heavy-chain-only antibodies changed how the field thought about antibody recognition.

Alongside conventional antibodies built from paired heavy and light chains, camelids also produce antibodies without light chains. The binding domain of these heavy-chain-only antibodies, the VHH, can fold independently and recognize an antigen on its own.

That discovery provided the essential natural proof behind the single-domain antibody:

One compact domain can function as a complete binding unit.

Camelid immunization has since become one of the most established routes to VHH discovery. It has produced exceptional binders and helped create an important class of research, diagnostic, and therapeutic molecules.

That success is real.

But the natural system that revealed the format was built to protect a camelid.

It was not built to produce a development-ready human therapeutic.

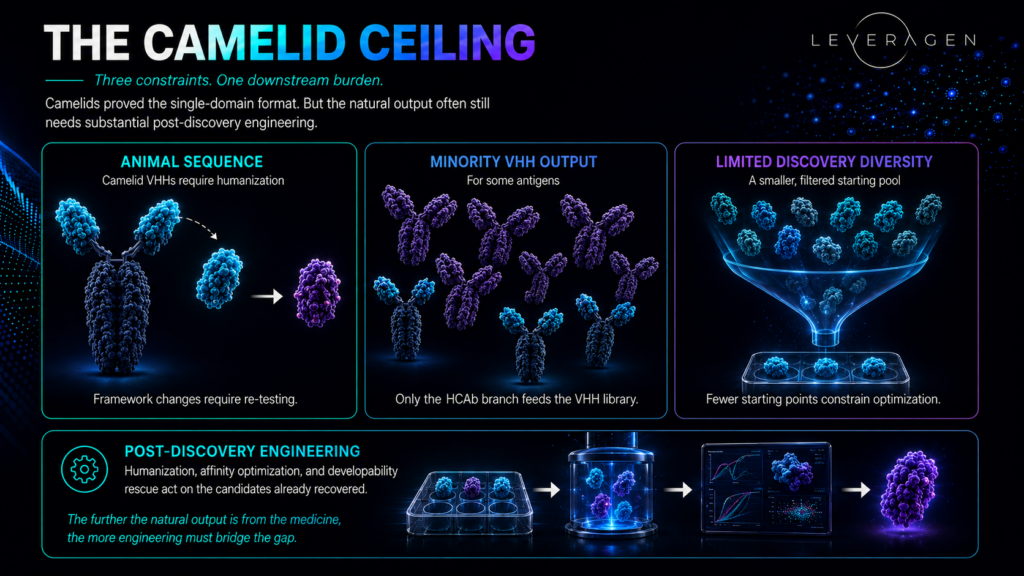
That distinction creates three separate constraints: the sequence begins as animal, the VHH-bearing response can be a minority output, and the effective diversity available to discovery is narrowed before screening is complete.

Each constraint matters on its own.

Together, they move a substantial part of molecule creation downstream.
The sequence begins as animal

A VHH recovered from a camelid carries a camelid sequence framework.

That framework is part of a molecule shaped by the animal’s immune system. It supports the binding loops, contributes to the domain’s structure, and helps determine properties such as stability, solubility, expression, and self-association.

For a human therapeutic, the recovered sequence typically cannot be treated as finished.

It often requires human-likeness assessment, humanization, or other sequence optimization to address potential immunogenicity and development concerns.

Humanization is not a cosmetic change.

The binding loops do not operate independently of the framework beneath them. Framework residues help position those loops and can influence the geometry of the antigen-binding surface.

Replacing camelid residues with human counterparts can therefore change more than the sequence label.

Affinity may shift. Stability may decline. Expression may change. Solubility may improve in one variant and worsen in another. A substitution intended to make the molecule more human-like may disturb the structural balance selected during the original immune response.

Every revised molecule must be tested again.

If its properties change, additional variants may need to be designed, expressed, and screened. Some camelid residues may need to be retained. Other positions may require compensating changes.

Humanization can be highly successful.

The limitation is not that a camelid VHH cannot become a suitable human therapeutic. It is that conversion introduces a new design cycle after discovery.

The binder found by the animal and the molecule advanced toward the clinic may not be the same sequence.

The VHH output can be the minority

Camelids do not produce VHH-bearing antibodies in isolation.

Their immune system generates conventional IgG1 antibodies alongside heavy-chain-only antibodies. Conventional antibodies recognize an antigen through a binding surface formed by paired heavy and light chains. Heavy-chain-only antibodies bind without light chains, using one VHH domain on each arm.

Both are complete antibody formats.

Only one supplies natural VHH sequences to a single-domain discovery campaign.

For some antigens, the heavy-chain-only response is strong and produces highly mature VHHs. For others, conventional antibodies take the larger share of the antigen-specific response.

The balance varies by species, animal, antigen, immunization schedule, sampling time, and assay.

A strong total response is not always a strong VHH response.

An immunized animal may generate abundant target-binding antibodies while only a smaller fraction belongs to the heavy-chain-only compartment.

If total antibody titer is used as the principal measure of success, that distinction can remain hidden until discovery begins.

The animal has responded.

But much of the response may be in a format the VHH campaign cannot use.

A minority VHH response does not mean the recovered domains will be weak. A smaller branch can still contain excellent binders.

It does mean that fewer VHH-producing lineages may be available to enter the campaign, even when the overall immune response appears robust.

The ceiling is not the quality any individual VHH can reach.

It is how much of the immune response enters the desired format in the first place.

Diversity narrows before the screen begins

The diversity of an immune response is larger than the diversity captured in a discovery library.

For camelid VHH discovery, the first narrowing occurs at the level of antibody format. Conventional heavy-light-chain antibodies are part of the animal’s response, but they do not contribute intact natural VHH binders.

Only the heavy-chain-only compartment is eligible.

The pool narrows again through biology. Some VHH lineages expand strongly. Others remain rare, disappear during selection, or never become sufficiently represented to be recovered.

The pool narrows again during discovery.

Sampling captures the response at a particular place and time. Amplification, cloning, microbial growth, display, and screening can favor some sequences over others. Rare lineages may be missed. Difficult-to-express domains may be underrepresented. Selection conditions may enrich what performs well in the assay rather than everything the immune system produced.

The final library is a filtered record of one compartment of the response.

This is not a claim that every camelid VHH library is small or uniform.

Camelids can produce substantial sequence diversity, unusual binding geometries, and exceptional domains.

The more precise issue is recovered discovery diversity: how many independent VHH lineages, epitopes, and development alternatives are actually present and accessible when screening begins.

A campaign may still find one strong candidate.

But a program rarely needs affinity alone.

If the leading VHH later shows poor stability, weak expression, nonspecific binding, an unfavorable epitope, or inadequate function, the value of the pool depends on the alternatives behind it.

A deeper pool offers more independent ways to solve the problem.

A narrower pool places more pressure on engineering the candidates already recovered.

The burden moves downstream

The three constraints converge after discovery.

The sequence may need to become more human-like.

Affinity or functional potency may need to improve.

Stability, solubility, expression, specificity, or other developability properties may need to be rescued.

These are familiar and often manageable tasks. Modern protein engineering can create remarkable improvements in a recovered molecule.

But the location of that work matters.

The natural output often becomes the beginning of a second discovery process.

Humanization changes the framework and requires revalidation. Affinity maturation creates and screens new variants. Developability engineering changes residues to address liabilities that may not have been visible during the original campaign.

Each intervention moves the molecule farther from the sequence selected in the animal.

The resulting candidate may be better suited to development.

It is also increasingly the product of deliberate post-discovery design.

This is what makes the engineering artificial: not ineffective or illegitimate, but performed outside the natural immune process.

Instead of mutation and selection occurring within the full biology of an immune response, variants are created and judged through chosen assays. Those assays can measure what a development team knows to test: binding, potency, expression, aggregation, stability, specificity, and other defined properties.

They cannot represent every biological pressure the molecule may encounter.

Engineering is powerful, but it is assay-bounded.

It can explore sequence space around the candidates in hand.

It cannot restore VHH lineages that never expanded in the animal. It cannot recover diversity lost during sampling or library construction. It cannot replay every path of mutation and selection that might have occurred in a more dedicated single-domain response.

Engineering can improve the starting point.

It cannot create the immune history that the starting point never received.

The problem is distance

None of these constraints invalidates camelid VHH discovery.

Camelids remain an important source of single-domain antibodies. Their best VHHs can combine high affinity, useful epitope recognition, compact size, and strong functional activity.

The issue is not whether a camelid can produce an exceptional molecule.

It is how much distance remains between the natural output and the intended medicine.

That distance has three dimensions.

The sequence is animal and often requires humanization.

The VHH-bearing branch may represent only a minority of the antigen-specific response.

The diversity available to discovery is narrowed by both immune biology and the process used to capture it.

The greater the combined distance, the more work moves downstream.

More variants must be designed. More properties must be retested. More tradeoffs must be managed after the original binder has already been selected.

The risk is not simply additional time.

Every change can alter the balance that made the original molecule attractive.

A humanizing substitution may affect affinity. An affinity-enhancing mutation may reduce stability. A solubility improvement may change function. A developability correction may require another round of screening.

The molecule can improve through this process.

But the path becomes one of serial repair and revalidation.

Format matters at the source

Camelids proved that a single domain could perform the complete binding function.

That discovery remains foundational.

But a natural source that reveals a format is not necessarily the ideal architecture for producing a human therapeutic in that format.

If the intended medicine is a human single-domain antibody, the strategic question is not only how to optimize a camelid VHH after discovery.

It is whether discovery can begin closer to the desired endpoint.

A format-native system would direct the immune response toward the single-domain architecture from the beginning. A human sequence framework would reduce the need for later conversion. A dedicated output would avoid dividing the antigen-specific response between usable and unusable antibody formats. Broader recovered diversity would provide more independent starting points before engineering begins.

That does not eliminate development work.

Affinity, function, specificity, stability, expression, and manufacturability still matter. No discovery architecture makes every candidate automatically suitable for a medicine.

It changes what the engineering is asked to do.

Instead of transforming an animal-derived output into the intended therapeutic format, engineering can focus on refining candidates already closer to that format.

Leveragen’s Singularity platform was built around that premise: a light-chain-free mouse in which a human VH repertoire matures in vivo without a parallel paired-antibody arm.

The central idea is simple.

The closer the natural discovery output is to the intended medicine, the less artificial engineering must bridge the gap after discovery.

Camelids proved the single-domain antibody.

The next generation of discovery systems must ask whether the immune response itself can be designed around it.

Format matters in the final medicine.

It also matters in the system that creates the binder.

The Single-Domain Antibody Is Not Just a Smaller Antibody

Why one compact binding domain can support a wider range of biologic designs

The conventional full-length IgG has long defined what an antibody medicine looks like.

It remains one of biotechnology’s most successful molecular formats, combining target recognition, bivalent binding, long serum exposure, Fc-mediated biology, and a well-established development path in a single molecule.

For many programs, that is exactly what the medicine needs.

But the antibody is no longer always the entire medicine.

It may be one recognition arm in a multispecific. It may direct a T cell through a CAR. It may carry a payload, be encoded by mRNA or AAV, or sit on a lipid nanoparticle to tell the vehicle where to go.

In these applications, the antibody is a component inside a larger therapeutic design.

That changes what the ideal component looks like.

In the first article of this series, The Discovery Engine Has Not Kept Up with the Medicine, we argued that the final modality should help determine the antibody format from the beginning. Single-domain antibodies are an important example of why that matters.

At approximately 12-15 kDa, a single-domain antibody, or sdAb, is about one-tenth the size of a conventional IgG. It is one independently functioning variable domain encoded by one compact sequence.

Its defining advantage is not simply that it is smaller.

It is that one domain can perform the complete binding function.

That architectural simplicity creates applications that larger or more assembly-dependent antibody formats do not always support as easily.

One domain changes the engineering problem

A full-length IgG depends on the expression and correct assembly of four polypeptide chains. A Fab is smaller, but its binding site still comes from paired heavy- and light-chain variable domains. An scFv places those two domains on one polypeptide but still relies on their orientation and a peptide linker.

Each format remains highly useful.

An sdAb starts from a different premise: the binder itself is one domain.

There is no light chain to pair and no VH-VL linker to optimize. One sequence corresponds to one binding unit.

That does not make every sdAb automatically stable, soluble, high-affinity, or developable. Those properties still need to be selected and tested.

But when a binder must be moved, fused, repeated, encoded, conjugated, or displayed, removing one layer of architectural dependency can matter.

Modularity: building complex biology from simple parts

The biology of a multispecific may be complicated.

The building blocks do not have to be.

sdAbs can be linked in tandem to create bispecific and multispecific constructs. They can be reordered, repeated to increase avidity, combined with Fc regions, or fused to enzymes, cytokines, receptors, and other functional modules.

Because each binding unit is self-contained, the design does not have to manage a separate light chain for every specificity. That can reduce pairing complexity and make more geometries practical to test.

Geometry still matters. Domain order, linker length, valency, epitope position, and spacing can determine whether a construct bridges two cells, clusters a receptor, blocks parallel pathways, or remains inactive until the right targets are present.

sdAbs do not answer those design questions automatically.

They provide a cleaner set of parts with which to explore them.

The same principle applies to multivalent formats. Repeating one domain can increase avidity against dense or repeating targets, while combining domains against different epitopes can create cooperative binding or reduce escape.

The molecule can become more complex without making each recognition unit more complex.

Cell therapy and immune engagement: a compact recognition domain

In a CAR, the antibody-derived binding domain is not a circulating drug.

It is the extracellular sensor of an engineered cell.

That sensor must be encoded within a larger receptor, expressed on the cell surface, and function in the geometry imposed by the CAR architecture. In many current designs, that role is performed by an scFv.

Single-domain antibodies provide another option.

Their compact architecture can simplify receptor construction and support tandem or multispecific CARs in which several recognition domains must fit within one genetic cassette. Fully human sdAbs can also provide human recognition domains without a separate humanization step.

The same modularity can support T-cell engagers and other immune-cell redirectors. One sdAb may recognize the disease target while another engages an immune receptor, with additional modules used to tune valency, half-life, or conditional activity.

Performance still depends on affinity, epitope, receptor spacing, antigen density, and signaling design.

The sdAb gives engineers a compact way to implement those biological choices.

Encoded biologics: when sequence length becomes a therapeutic property

The value of a one-domain binder becomes especially clear when the medicine is delivered as genetic information.

A conventional antibody requires separate heavy and light chains to be expressed in the same cell and assembled correctly. An sdAb can be encoded as one functional polypeptide from one compact coding sequence.

For mRNA medicines, that can simplify construct design and make it easier to encode multivalent or multispecific proteins within one open reading frame.

For AAV and other viral vectors, sequence space is limited. Every nucleotide used for the therapeutic protein competes with promoters, signal peptides, regulatory elements, and other components needed for expression.

A compact sdAb leaves more room for the rest of the cassette.

In these settings, sequence length is not merely convenient.

It is part of the therapeutic architecture.

Conjugates and targeted delivery: the antibody as an address

In an ADC or targeted delivery system, the binding domain has a defined job: recognize the destination and bring something else with it.

sdAbs can serve as compact targeting elements in drug conjugates, radionuclide conjugates, protein fusions, lipid nanoparticles, viral particles, and other delivery vehicles.

Their well-defined scaffold can support site-specific conjugation and controlled orientation when the appropriate residues, tags, or linkers are engineered into the molecule. Their limited footprint may also allow targeting ligands to be displayed without adding the mass of a full-length antibody.

This is particularly relevant for lipid nanoparticles.

Decorating an LNP with a cell-selective sdAb can give the particle an address beyond its default biodistribution. In principle, the same vehicle can be redirected toward a tumor antigen, an immune-cell subset, or another tissue marker by changing the targeting domain.

But a ligand does not solve delivery by itself. Density, orientation, circulation, protein corona formation, cellular uptake, and endosomal escape all influence whether the payload becomes active in the intended cell.

The sdAb provides the recognition module.

The vehicle still has to complete the journey.

Difficult targets: access depends on geometry

Affinity is not the only determinant of what an antibody can do.

The geometry of the binding surface also matters.

Conventional antibodies recognize antigens through a broad interface formed by paired heavy- and light-chain variable domains. That architecture is extraordinarily effective across many targets.

Some useful epitopes, however, are recessed, transient, conformational, or located close to a membrane.

The compact paratope of an sdAb can approach these surfaces differently. In several single-domain scaffolds, extended complementarity-determining loops can reach into clefts, pockets, and other constrained regions that may be difficult for a larger paired surface to engage.

This has made sdAbs valuable tools for conformationally dynamic membrane proteins, including GPCRs, and promising binders for ion channels and other difficult target classes. They can recognize extracellular loops, stabilize defined receptor states, or bind pore-adjacent and cryptic epitopes.

The advantage is not that every sdAb reaches every hidden site.

It is that the format expands the accessible epitope space.

When epitope determines mechanism, that can change the biology available to a program.

Tissue access: smaller molecules move differently

A 12-15 kDa binder distributes differently from a 150 kDa IgG.

sdAbs can diffuse rapidly through dense tissue and may penetrate solid tumors more evenly than larger antibodies, which can be slowed by size and high-affinity capture near the tumor periphery.

That can support therapeutic delivery, as well as support imaging.

Rapid tissue entry followed by fast clearance from the blood can create high target-to-background contrast sooner than a long-circulating IgG. The same pharmacokinetic profile that limits chronic systemic exposure can therefore benefit diagnostics, image-guided surgery, or short-duration targeting.

The blood-brain barrier requires more nuance.

Small size alone does not make an antibody brain-penetrant. Most proteins still require an active transport mechanism to cross efficiently.

But sdAbs can serve as compact brain-shuttle modules when selected against receptors that undergo transcytosis. The shuttle can be fused to a larger cargo and used to increase its central nervous system exposure.

The opportunity is not passive passage because the molecule is small.

It is the ability to build a compact transport module into a larger medicine.

Intracellular applications: reaching beyond the cell surface

Most therapeutic antibodies act outside the cell because conventional antibody fragments often do not fold or remain soluble in the reducing cytoplasm.

Some single-domain antibodies can function there.

When selected or engineered for intracellular stability, an sdAb can be expressed as an intrabody and bind a target in the compartment where it operates. It can inhibit an interaction, trap a protein in a location, report on a conformational state, or be fused to an effector that changes the target’s fate.

That creates potential access to transcription factors, signaling proteins, oncoproteins, viral proteins, and other intracellular targets that are difficult to approach with extracellular antibodies.

This capability is sequence-dependent. Intracellular candidates must be screened under the conditions in which they are expected to function.

But one genetically encoded domain provides a practical architecture for that search.

The addressable target space can move from the cell surface toward the intracellular proteome.

New routes of administration: bringing the binder to the disease

Full-length antibodies are usually administered by injection.

Their size, formulation requirements, and poor permeability across epithelial barriers make alternative routes difficult.

The compactness and robustness of some sdAbs can support local delivery strategies that are less practical for conventional antibodies.

Inhaled sdAbs can be delivered directly to the respiratory tract. Topical or ocular formats can place the binder at a disease surface. Orally administered sdAbs are also being developed for local action in the gastrointestinal tract, where formulation and protease stability can help the molecule reach intestinal targets.

Oral systemic delivery remains a much harder problem. An sdAb still faces gastric acid, digestive enzymes, mucus, and the epithelial barrier.

The opportunity is not that every sdAb can become a pill.

It is that the format can make selected local routes realistic enough to engineer.

The tradeoffs are part of the format

sdAbs are not miniature IgGs with every IgG property preserved.

They generally lack Fc-mediated functions unless an Fc or another immune-engaging module is added. Their small size can produce rapid renal clearance and short systemic half-life. A monovalent sdAb may need multimerization for the required avidity. Some sequences will need further engineering for solubility, stability, expression, or immunogenicity.

These are real constraints.

They are also design variables.

Half-life can be extended through Fc fusion, albumin binding, PEGylation, multimerization, or other approaches. Effector function can be added when the mechanism requires it. Rapid clearance can be retained for imaging or local therapy rather than engineered away.

The objective is not to make an sdAb behave exactly like an IgG.

It is to add the functions the application needs while preserving the advantages of the smaller binding unit.

Versatility is the point

The future of biologics will continue to depend on conventional full-length antibodies.

When a program needs long exposure, Fc biology, bivalent binding, and a deeply established development path, the IgG remains an exceptional format. Fabs, scFvs, and other fragments will remain important as well.

Single-domain antibodies occupy a distinct position within that landscape.

They offer one binding domain, one compact sequence, and a modular unit that can be linked, repeated, encoded, conjugated, displayed, transported, or expressed inside a cell.

That versatility supports a remarkable range of applications: multispecific antibodies, CARs and immune engagers, ADCs, mRNA and AAV medicines, targeted nanoparticles, difficult membrane proteins, imaging, brain delivery, intrabodies, and local administration.

The question is not whether the sdAb is universally better than every other antibody format.

It is whether its architecture is better matched to the medicine being built.

In the next article in this series, we will turn from the format to the discovery system and examine how Leveragen’s Singularity Suite is designed to generate sdAbs directly in vivo.

The molecule is smaller. What it enables is not.

The Discovery Engine Has Not Kept Up with the Medicine

Why next-generation biologics need format-native antibody discovery

For decades, antibody discovery was optimized around one extraordinarily successful output: the conventional full-length IgG.

The classic four-chain antibody — two heavy chains and two light chains — remains one of the most validated molecular formats in biotech. It has produced some of the most important therapeutics in modern medicine, such as Keytruda and Rituxan for cancer treatment and Humira and Dupixent for chronic inflammation. And it continues to be the right format for many programs.

But the antibody field is no longer a one-format field.

Today’s therapeutic frontier includes bispecifics, multispecifics, ADCs, T-cell engagers, CAR architectures, encoded antibodies, targeted delivery systems, and other complex biologic modalities. In these settings, the antibody is not always the final therapeutic format by itself. Increasingly, it functions as a modular component inside a larger design.

It may serve as a targeting domain.
It may serve as a payload carrier.
It may define the recognition domain of an engineered cell therapy.
It may be genetically encoded.
It may need to be fused, conjugated, multimerized, paired, reformatted, or combined with another binding specificity.

That shift changes what antibody discovery needs to deliver.

The problem is not that conventional antibodies have failed. They have transformed medicine. The problem is that many discovery systems were built for an era in which the default output was a full-length IgG, while many of today’s modalities require antibody building blocks with different architectural constraints.

The discovery engine has not fully kept up with the medicine.

The bottleneck: discovery format versus final modality

In many antibody programs, discovery begins in one format and development ends in another.

A team may identify a promising binder as a conventional heavy-chain/light-chain antibody, then later reformat it into a bispecific, multispecific, ADC component, CAR recognition domain, encoded biologic, or delivery module. That approach can work, and the field has developed many sophisticated engineering strategies to support it.

However, it also creates a recurring bottleneck: the molecule is often discovered before the final format has been fully imposed on it.

As a result, format liabilities can appear late.

A binder that looks promising in one architecture may behave differently after reformatting. It may lose functional geometry. It may express poorly. It may aggregate. It may create purification challenges. It may introduce manufacturability concerns. It may require extensive engineering before it becomes compatible with the modality the program actually needs.

In other words, the issue is not only binding. It is also whether the binder was discovered in an architecture aligned with its eventual use.

Chain pairing is one visible example

Bispecific antibodies make this problem especially clear.

In a conventional antibody, biological function depends on precise pairing between heavy and light chains. In a bispecific or multispecific setting, multiple chains may need to be expressed together and assembled correctly. Without the correct architecture or engineering strategy, the desired molecule can become only one species among several possible pairing outcomes.

The consequences are familiar to anyone working in the field: reduced yield of the intended molecule, additional purification burden, product heterogeneity, stability concerns, and time spent managing molecular assembly rather than advancing biology.

The industry has developed many ways to address this problem downstream. Those strategies have enabled important progress and will continue to matter.

But the deeper question is strategic:

Should format compatibility always be solved after discovery?

Or should discovery systems be designed to generate antibody building blocks closer to the format the modality actually requires?

At Leveragen, our answer is clear:

The modality should help determine the discovery format from the start.

Toward format-native antibody discovery

Leveragen’s models have been built around the idea that antibody discovery should not be limited to a single default output.

Instead of discovering conventional antibodies first and asking downstream teams to convert them later, Leveragen develops in vivo platforms designed to generate different antibody architectures natively.

We call this approach format-native in vivo antibody discovery.

The premise is simple: if the final modality requires a single-domain binder, discover single-domain antibodies from the beginning. If the final modality requires bispecific compatibility, discover binders in a common light-chain context from the beginning. If the biology calls for a conventional full-length IgG, preserve that proven format and discover accordingly.

The goal is not to declare one antibody format superior to all others. Rather, the goal is to match the discovery system to the modality.

That is why Leveragen has built three complementary in vivo antibody discovery platforms: SingularityUniversality, and Infinity.

Singularity: single-domain antibodies for modular biologics

The most direct way to remove heavy-light chain pairing dependency is to remove the light chain entirely.

Leveragen’s Singularity Suite is designed for the discovery of single-domain antibodies: compact, single-chain binders that can serve as modular components across multiple biologic formats.

Single-domain antibodies are attractive because their architecture is simple. They do not require heavy-light chain pairing. Their compact size can make them well suited for multivalent and multispecific formatting, genetic fusion, payload-conjugation designs, CAR recognition domains, encoded antibody approaches, and targeted delivery applications.

For many next-generation modalities, that compact architecture is not just convenient; it can be central to the design.

A single-domain binder can be used as a module. It can be combined with other domains. It can be encoded as a single genetic unit. It can access epitopes that may be difficult for larger antibody formats to reach. It can support formats where size, geometry, and modularity are critical.

Leveragen’s Singularity platform is designed to generate these binders in vivo rather than forcing conventional antibodies into single-domain-like applications after discovery.

That distinction matters.

A binder discovered in the intended architecture has already been shaped by the constraints of that architecture. The starting point is closer to the final modality. That can reduce the need for late-stage reformatting or rescue engineering and give development teams a cleaner path from discovery to design.

Universality: common light-chain antibodies for bispecific development

Not every program needs a single-domain antibody.

Many programs still benefit from an antibody architecture closer to a conventional IgG, but they also need compatibility with bispecific development and scalable manufacturing.

This is where Leveragen’s Universality Suite comes in.

The common light-chain strategy addresses one of the central challenges in bispecific development: light-chain mispairing. In a conventional bispecific setting, two different heavy chains may each require their own light-chain partner. When these chains are expressed together, multiple assembly products can result.

A common light-chain approach reduces that complexity by placing diverse heavy-chain binders into a shared light-chain context.

This does not mean every downstream engineering challenge disappears. Heavy-chain heterodimerization, manufacturability, developability, stability, and format-specific design still matter. But by reducing light-chain combinatorial complexity, common light-chain discovery can create a more direct path toward bispecific assembly.

The key is to solve part of the format problem upstream.

Instead of discovering two antibodies and later asking whether their light chains can be made compatible, Universality is designed to discover heavy-chain diversity in a common light-chain framework from the beginning.

That makes the resulting binders more naturally aligned with bispecific development.

For teams building bispecifics, this can shift the starting point. The question becomes less about whether two independently discovered antibodies can be forced into a compatible architecture, and more about how to select the best heavy-chain binders within a format already designed for bispecific assembly.

Infinity: full-length antibodies when the proven format is right

The rise of new modalities does not diminish the importance of conventional antibodies.

In many programs, the full-length IgG remains exactly the right answer.

Some therapeutic mechanisms benefit from the pharmacokinetics, effector functions, valency, stability, and established developability profile of a conventional antibody. Many diagnostic and veterinary applications also continue to rely on full-length antibody formats. The decades of biological, regulatory, and manufacturing experience behind conventional antibodies remain highly valuable.

That is the role of Leveragen’s Infinity Suite.

Infinity is designed for discovery of conventional full-length antibodies when the biology and application call for the classic format. It preserves the value of intact heavy- and light-chain antibody discovery while supporting broad applications across therapeutics and diagnostics.

This is important to the overall Leveragen thesis.

The future is not single-domain antibodies instead of conventional antibodies.
It is not common light-chain bispecifics instead of full-length IgGs.
It is not one platform replacing every other platform.

The future is format choice.

Different modalities require different antibody architectures. A discovery engine should be able to produce the right one.

The shift: from target-first to target-plus-format discovery

Historically, antibody discovery has often been framed as a target-first process: identify the antigen, generate binders, then engineer the molecule into the therapeutic format needed later.

That model will remain useful in many cases. But as biologics become more architecturally complex, discovery needs to become more format-aware from the beginning.

For next-generation modalities, the key question is not only:

Does this antibody bind the target?

It is also:

Does this antibody work in the format the modality requires?

That second question is increasingly decisive.

A binder for a CAR recognition domain may need different properties from a binder intended to become a full-length IgG. A binder for an ADC may need different internalization behavior and conjugation compatibility. A binder for a bispecific may need pairing compatibility and functional geometry. A binder for encoded delivery may need compact genetic architecture. A binder for a multispecific may need to behave well as one domain inside a larger molecule.

These are not downstream details; they are discovery-relevant constraints.

That is why Leveragen’s platform strategy is built around matching antibody architecture to modality requirements earlier in the process.

A discovery engine for a multi-format field

The antibody field has expanded beyond a single dominant molecular output.

That expansion is a sign of progress. Biologics are becoming more programmable, more modular, and more capable of addressing complex disease biology. But the discovery infrastructure must evolve with that progress.

If the medicine becomes multi-format, discovery must become multi-format too.

Leveragen’s approach is built around three complementary outputs:

Singularity for single-domain antibodies when the modality benefits from compact, single-chain, modular binders.

Universality for common light-chain antibodies when bispecific development requires reduced light-chain pairing complexity.

Infinity for conventional full-length antibodies when the proven IgG architecture remains the right format.

Together, these platforms reflect a simple proposition:

The right antibody building block should be discovered in the right format, in vivo, from the start.

This is the gap Leveragen is built to close.

The next generation of biologics will not be built from one antibody format alone. It will require single-domain antibodies, common light-chain antibodies, conventional full-length antibodies, and other architectures matched to specific therapeutic needs.

The modality should determine the format. And the discovery engine should be able to keep up.

What this series will cover

This essay introduces the central idea: next-generation biologics need format-native antibody discovery.

In the coming posts, we will examine each part of this platform strategy in more detail:

Why single-domain antibodies are becoming increasingly important as modular biologic building blocks.

Why common light-chain discovery can change the starting point for bispecific development.

Why conventional full-length antibodies remain essential when the biology calls for the classic IgG format.

And why in vivo discovery continues to matter in an era increasingly shaped by display technologies, synthetic libraries, computational design, and AI.

The future of antibody therapeutics is not one format; it is the right format for the right modality.